Aegis provides affordable, low power Wi-Fi sensor network solutions that make a ubiquitous sensing world come alive and a full line of controlled temperature cabinets. Laboratory, Pharmacy and Chromatography refrigerators and freezers which are designed to meet the dimensional and temperature requirements of the scientific community.
Why Choose ENT Medical Equipment for Your Clinic?
Choosing the right Ent Medical Equipment can shape every consultation, from the first patient greeting to post-treatment follow-up. In a busy clinic, equipment must support clear examination, safe procedures, and confident clinical decisions. A reliable otoscope, endoscope, suction unit, or treatment chair is more than a purchase. It becomes part of the daily workflow. Small details matter. Smooth controls reduce interruptions. Bright, accurate lighting reveals subtle tissue changes. Comfortable positioning can ease anxiety for both patients and clinicians.
Experienced ENT teams usually assess equipment through practical use, not attractive specifications alone. They consider image quality, sterilization requirements, ergonomic design, repair access, and staff training. A device may perform well during a demonstration yet feel awkward after several hours of consultations. That assumption can fail. Clinicians should review manufacturer documentation, applicable safety standards, warranty terms, and service records before committing. Independent clinical guidance can also help identify features that genuinely improve diagnosis or treatment.
No device is perfect. Budget limits, room size, patient volume, and staff experience all influence the best choice. A compact examination system may suit a small practice, while a hospital department may need advanced imaging and stronger integration. Careful evaluation reduces wasted investment and supports consistent patient care. Still, technology cannot replace clinical judgment. It should make good practice easier, not pretend to make decisions alone. With documented maintenance, responsible training, and honest feedback from users, Ent Medical Equipment can become a dependable foundation for a safer, more efficient ENT clinic.
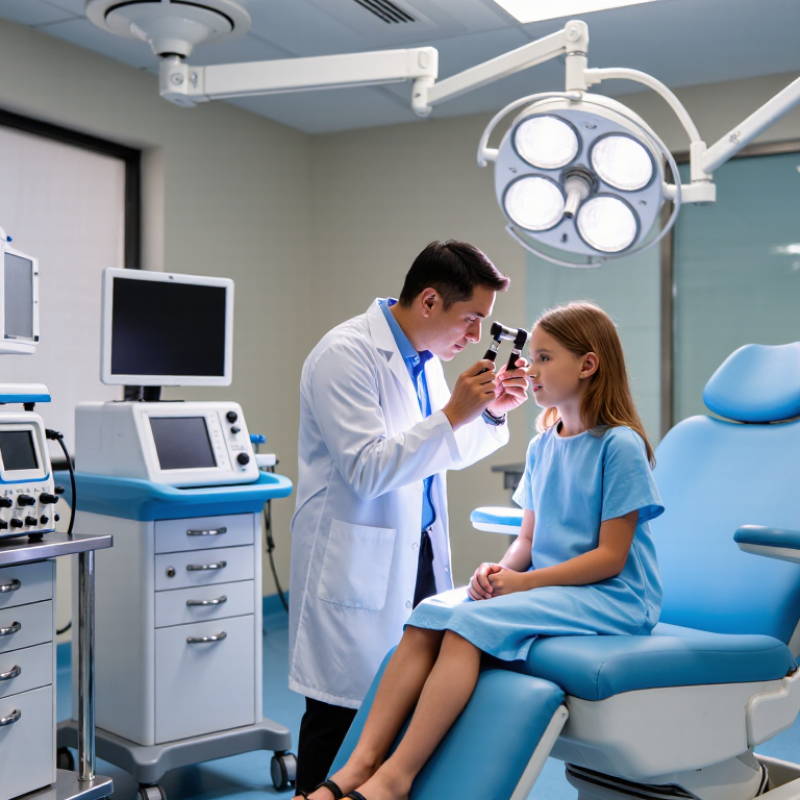
Table of Contents [Hide]
Assessing ENT Demand: WHO Reports 1.5 Billion People With Hearing Loss
Why Choose ENT Medical Equipment for Your Clinic?
The World Health Organization reports that more than 1.5 billion people live with hearing loss worldwide. Its World Report on Hearing also projects that this number could reach 2.5 billion by 2050. Around 430 million people currently need rehabilitation services, including hearing assessment and communication support. These figures show significant demand for ENT services, even in clinics that do not consider themselves hearing specialists.
The number is powerful, but it is not a complete local forecast. Patient age, occupational noise, childhood infections, and access to healthcare can change regional demand. A clinic needs equipment that supports accurate, repeatable examinations. A calibrated audiometer can measure hearing thresholds, while tympanometry helps assess middle-ear function. An otoscope with clear illumination can reveal wax, inflammation, or visible membrane changes. Small details matter.
WHO emphasizes that timely identification can reduce the impact of hearing loss. In practice, this means a quiet testing room, documented calibration, and trained staff are as important as the devices themselves. Infection-control procedures also protect patients during repeated examinations. The Global Burden of Disease Study 2019, published in The Lancet, identified hearing loss as one of the leading causes of years lived with disability. That evidence supports wider diagnostic capacity, but it does not remove uncertainty. Some clinics may overinvest in advanced tools while overlooking basic workflow problems. Equipment should match patient volume, clinical skills, referral pathways, and maintenance resources. Calibration records must stay current. Mistakes still happen.
Contact Us
Phone
Toll Free: 1-800-796-2344
OH: 513 204 5837
PA: 215-281-9180
support@site_367d8ed1-a876-442b-a1d7-b1fc1cf4ed19